
van der Spoel labb
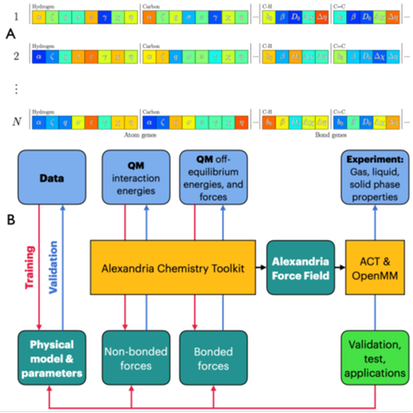
David van der Spoel’s forskargrupp arbetar med maskininlärning för att framställa kraftfält för molekylsimulering. De använder stora databaser med kvantkemiska beräkningar på molekyler (se bild) och har implementerat en mjukvara kallad Alexandria Chemistry Toolkit (ACT) för att automatiskt ta fram kraftfält. Mjukvaran gör det möjligt att jamföra hur pass bra olika matematiska ekvationer kan reproducera referensdatat. Gruppen har publicerat en hel del artiklar om metoderna, till exempel en om ädelgaser där vi visar att våra modeller är tillräckligt noggranna för att reproducera smältpunkter på ädelgaskristaller (J. Chem. Theory Comput 20 (2024) 2362–2376). En annan artikel beskriver upptäckten av så-kallade sigmahål i vatten, längsmed OH bindningarna (J. Phys. Chem. Letters 158 (2024) 9974). Nu senast har vi använt ACT för att ifrågasätta en populär metod för att bestämma partiella laddningar i kraftfält. Inom kort hoppas vi kunna färdigställa ett komplett kraftfält för proteiner i lösning.
Gruppmedlemmar
Publikationer
-
LacI strikes a balance between stability and inducibility
Ingår i Nucleic Acids Research, 2026
- DOI för LacI strikes a balance between stability and inducibility
- Ladda ner fulltext (pdf) av LacI strikes a balance between stability and inducibility
-
plotXVG: Batch Generation of Publication-Quality Graphs from GROMACS Output
Ingår i Journal of Chemical Information and Modeling, s. 2955-2959, 2026
- DOI för plotXVG: Batch Generation of Publication-Quality Graphs from GROMACS Output
- Ladda ner fulltext (pdf) av plotXVG: Batch Generation of Publication-Quality Graphs from GROMACS Output
-
Beyond Partitioning: Using Force Field Science to Evaluate Electrostatics Models
Ingår i Journal of Chemical Theory and Computation, s. 2220-2231, 2026
- DOI för Beyond Partitioning: Using Force Field Science to Evaluate Electrostatics Models
- Ladda ner fulltext (pdf) av Beyond Partitioning: Using Force Field Science to Evaluate Electrostatics Models
-
Quantitative evaluation of anharmonic bond potentials for molecular simulations
Ingår i Digital Discovery, s. 824-830, 2025
- DOI för Quantitative evaluation of anharmonic bond potentials for molecular simulations
- Ladda ner fulltext (pdf) av Quantitative evaluation of anharmonic bond potentials for molecular simulations
-
Point + Gaussian charge model for electrostatic interactions derived by machine learning
Ingår i Physical Chemistry, Chemical Physics - PCCP, s. 13817-13820, 2025
- DOI för Point + Gaussian charge model for electrostatic interactions derived by machine learning
- Ladda ner fulltext (pdf) av Point + Gaussian charge model for electrostatic interactions derived by machine learning
-
The need to implement FAIR principles in biomolecular simulations
Ingår i Nature Methods, s. 641-645, 2025
-
Evolutionary machine learning of physics-based force fields in high-dimensional parameter-space
Ingår i Digital Discovery, s. 1925-1935, 2025
- DOI för Evolutionary machine learning of physics-based force fields in high-dimensional parameter-space
- Ladda ner fulltext (pdf) av Evolutionary machine learning of physics-based force fields in high-dimensional parameter-space
-
Bayesian three-point water models
Ingår i npj Computational Materials, 2025
- DOI för Bayesian three-point water models
- Ladda ner fulltext (pdf) av Bayesian three-point water models
-
Ingår i Journal of Chemical Theory and Computation, s. 2362-2376, 2024
- DOI för Impact of Combination Rules, Level of Theory, and Potential Function on the Modeling of Gas- and Condensed-Phase Properties of Noble Gases
- Ladda ner fulltext (pdf) av Impact of Combination Rules, Level of Theory, and Potential Function on the Modeling of Gas- and Condensed-Phase Properties of Noble Gases
-
Martini on the Rocks: Can a Coarse-Grained Force Field Model Crystals?
Ingår i The Journal of Physical Chemistry Letters, s. 1079-1088, 2024
- DOI för Martini on the Rocks: Can a Coarse-Grained Force Field Model Crystals?
- Ladda ner fulltext (pdf) av Martini on the Rocks: Can a Coarse-Grained Force Field Model Crystals?
-
Assessment of Climate Impact of Sustainable Forestry Based on Landscape Structure
Ingår i Forests, 2024
- DOI för Assessment of Climate Impact of Sustainable Forestry Based on Landscape Structure
- Ladda ner fulltext (pdf) av Assessment of Climate Impact of Sustainable Forestry Based on Landscape Structure
-
Ingår i The Journal of Physical Chemistry Letters, s. 9974-9978, 2024
- DOI för Quantification of Anisotropy in Exchange and Dispersion Interactions: A Simple Model for Physics-Based Force Fields
- Ladda ner fulltext (pdf) av Quantification of Anisotropy in Exchange and Dispersion Interactions: A Simple Model for Physics-Based Force Fields
-
Simulations of Amyloid-Forming Peptides in the Crystal State
Ingår i The Protein Journal, s. 192-204, 2023
- DOI för Simulations of Amyloid-Forming Peptides in the Crystal State
- Ladda ner fulltext (pdf) av Simulations of Amyloid-Forming Peptides in the Crystal State
-
Can molecular dynamics be used to simulate biomolecular recognition?
Ingår i Journal of Chemical Physics, 2023
-
Probing Phase Transitions in Organic Crystals Using Atomistic MD Simulations
Ingår i ACS Physical Chemistry Au, s. 84-93, 2023
- DOI för Probing Phase Transitions in Organic Crystals Using Atomistic MD Simulations
- Ladda ner fulltext (pdf) av Probing Phase Transitions in Organic Crystals Using Atomistic MD Simulations
-
Binding Networks Identify Targetable Protein Pockets for Mechanism-Based Drug Design
Ingår i International Journal of Molecular Sciences, 2022
- DOI för Binding Networks Identify Targetable Protein Pockets for Mechanism-Based Drug Design
- Ladda ner fulltext (pdf) av Binding Networks Identify Targetable Protein Pockets for Mechanism-Based Drug Design
-
Ingår i Journal of Chemical Physics, 2021
-
Accurate absolute free energies for ligand-protein binding based on non-equilibrium approaches
Ingår i Communications Chemistry, 2021
- DOI för Accurate absolute free energies for ligand-protein binding based on non-equilibrium approaches
- Ladda ner fulltext (pdf) av Accurate absolute free energies for ligand-protein binding based on non-equilibrium approaches
-
Microscopic origins of conductivity in molten salts unraveled by computer simulations
Ingår i Communications Chemistry, 2021
- DOI för Microscopic origins of conductivity in molten salts unraveled by computer simulations
- Ladda ner fulltext (pdf) av Microscopic origins of conductivity in molten salts unraveled by computer simulations
-
Ingår i Journal of Chemical Physics, 2021
-
Editorial overview: Theory and simulation and their new friends
Ingår i Current opinion in structural biology, 2021
-
NMR Refinement and Peptide Folding Using the GROMACS Software
Ingår i Journal of Biomolecular NMR, s. 143-149, 2021
- DOI för NMR Refinement and Peptide Folding Using the GROMACS Software
- Ladda ner fulltext (pdf) av NMR Refinement and Peptide Folding Using the GROMACS Software
-
A potential for molecular simulation of compounds with linear moieties
Ingår i Journal of Chemical Physics, 2020
- DOI för A potential for molecular simulation of compounds with linear moieties
- Ladda ner fulltext (pdf) av A potential for molecular simulation of compounds with linear moieties
-
Toward a Computational Ecotoxicity Assay
Ingår i Journal of Chemical Information and Modeling, s. 3792-3803, 2020
-
Propagation of uncertainty in physicochemical data to force field predictions
Ingår i Physical Review Research, 2020
- DOI för Propagation of uncertainty in physicochemical data to force field predictions
- Ladda ner fulltext (pdf) av Propagation of uncertainty in physicochemical data to force field predictions
-
An Intuitively Understandable Quality Measure for Theoretical Vibrational Spectra
Ingår i The Journal of Physical Chemistry Letters, s. 5471-5475, 2020
- DOI för An Intuitively Understandable Quality Measure for Theoretical Vibrational Spectra
- Ladda ner fulltext (pdf) av An Intuitively Understandable Quality Measure for Theoretical Vibrational Spectra
-
Making Soup: Preparing and Validating Models of the Bacterial Cytoplasm for Molecular Simulation
Ingår i Journal of Chemical Information and Modeling, s. 322-331, 2020
-
Theoretical Infrared Spectra: Quantitative Similarity Measures and Force Fields
Ingår i Journal of Chemical Theory and Computation, s. 3307-3315, 2020
- DOI för Theoretical Infrared Spectra: Quantitative Similarity Measures and Force Fields
- Ladda ner fulltext (pdf) av Theoretical Infrared Spectra: Quantitative Similarity Measures and Force Fields
-
Role of Host-Guest Charge Transfer in Cyclodextrin Complexation: A Computational Study
Ingår i The Journal of Physical Chemistry C, s. 17745-17756, 2019
-
Molten alkali halides - temperature dependence of structure, dynamics and thermodynamics
Ingår i Physical Chemistry, Chemical Physics - PCCP, s. 18516-18524, 2019
-
Direct Link between Structure, Dynamics, and Thermodynamics in Molten Salts
Ingår i The Journal of Physical Chemistry C, s. 25596-25602, 2019
-
Systematically improved melting point prediction: a detailed physical simulation model is required
Ingår i Chemical Communications, s. 12044-12047, 2019
- DOI för Systematically improved melting point prediction: a detailed physical simulation model is required
- Ladda ner fulltext (pdf) av Systematically improved melting point prediction: a detailed physical simulation model is required
-
Prediction of Partition Coefficients of Environmental Toxins Using Computational Chemistry Methods
Ingår i ACS Omega, s. 13772-13781, 2019
- DOI för Prediction of Partition Coefficients of Environmental Toxins Using Computational Chemistry Methods
- Ladda ner fulltext (pdf) av Prediction of Partition Coefficients of Environmental Toxins Using Computational Chemistry Methods
-
Ingår i PloS Computational Biology, 2019
- DOI för Ten simple rules on how to create open access and reproducible molecular simulations of biological systems
- Ladda ner fulltext (pdf) av Ten simple rules on how to create open access and reproducible molecular simulations of biological systems
-
Rotational and Translational Diffusion of Proteins as a Function of Concentration
Ingår i ACS Omega, s. 20654-20664, 2019
- DOI för Rotational and Translational Diffusion of Proteins as a Function of Concentration
- Ladda ner fulltext (pdf) av Rotational and Translational Diffusion of Proteins as a Function of Concentration
-
Transient isomers in the photodissociation of bromoiodomethane
Ingår i Journal of Chemical Physics, 2018
-
Force Field Benchmark of Amino Acids: I. Hydration and Diffusion in Different Water Models
Ingår i Journal of Chemical Information and Modeling, s. 1037-1052, 2018
-
Statistical efficiency of methods for computing free energy of hydration
Ingår i Journal of Chemical Physics, 2018
- DOI för Statistical efficiency of methods for computing free energy of hydration
- Ladda ner fulltext (pdf) av Statistical efficiency of methods for computing free energy of hydration
-
Small Molecule Thermochemistry: A Tool for Empirical Force Field Development
Ingår i Journal of Physical Chemistry A, s. 8982-8988, 2018
-
Ingår i Scientific Data, 2018
- DOI för The Alexandria library, a quantum-chemical database of molecular properties for force field development
- Ladda ner fulltext (pdf) av The Alexandria library, a quantum-chemical database of molecular properties for force field development
-
Phase-Transferable Force Field for Alkali Halides
Ingår i Journal of Chemical Theory and Computation, s. 5933-5948, 2018
-
Ingår i Journal of Chemical Theory and Computation, s. 5553-5566, 2018
-
Impact of Dispersion Coefficient on Simulations of Proteins and Organic Liquids
Ingår i Journal of Physical Chemistry B, s. 8018-8027, 2018
-
Influence of Na+ and Mg2+ ions on RNA structures studied with molecular dynamics simulations
Ingår i Nucleic Acids Research, s. 4872-4882, 2018
- DOI för Influence of Na+ and Mg2+ ions on RNA structures studied with molecular dynamics simulations
- Ladda ner fulltext (pdf) av Influence of Na+ and Mg2+ ions on RNA structures studied with molecular dynamics simulations
-
Ingår i Journal of Chemical Theory and Computation, s. 1034-1043, 2017
-
Mechanistic Insights into Autoinhibition of the Oncogenic Chromatin Remodeler ALC1
Ingår i Molecular Cell, s. 847-8590000000, 2017
- DOI för Mechanistic Insights into Autoinhibition of the Oncogenic Chromatin Remodeler ALC1
- Ladda ner fulltext (pdf) av Mechanistic Insights into Autoinhibition of the Oncogenic Chromatin Remodeler ALC1
-
Systematic exploration of multiple drug binding sites
Ingår i Journal of Cheminformatics, 2017
- DOI för Systematic exploration of multiple drug binding sites
- Ladda ner fulltext (pdf) av Systematic exploration of multiple drug binding sites
-
Ingår i The Journal of Physical Chemistry Letters, s. 2705-2712, 2017
-
Ingår i Journal of Physics, 2017
-
Ingår i Journal of Chemical Physics, 2016
- DOI för Large-scale calculations of gas phase thermochemistry: Enthalpy of formation, standard entropy, and heat capacity
- Ladda ner fulltext (pdf) av Large-scale calculations of gas phase thermochemistry: Enthalpy of formation, standard entropy, and heat capacity
-
Exploration of Interfacial Hydration Networks of Target Ligand Complexes
Ingår i Journal of Chemical Information and Modeling, s. 148-158, 2016
-
Ingår i Journal of Chemical Information and Modeling, s. 2080-2092, 2016
- DOI för Evaluation of Generalized Born Models for Large Scale Affinity Prediction of Cyclodextrin Host-Guest Complexes
- Ladda ner fulltext (pdf) av Evaluation of Generalized Born Models for Large Scale Affinity Prediction of Cyclodextrin Host-Guest Complexes
-
Binding of Pollutants to Biomolecules: A Simulation Study
Ingår i Chemical Research in Toxicology, s. 1679-1688, 2016
- DOI för Binding of Pollutants to Biomolecules: A Simulation Study
- Ladda ner fulltext (pdf) av Binding of Pollutants to Biomolecules: A Simulation Study
-
Ingår i Journal of Chemical Physics, 2016
-
Atomistic Simulation of Protein Encapsulation in Metal-Organic Frameworks
Ingår i Journal of Physical Chemistry B, s. 477-484, 2016
-
Ingår i Journal of Chemical Theory and Computation, s. 5103-5113, 2015
-
Properties of Organic Liquids when Simulated with Long-Range Lennard-Jones Interactions
Ingår i Journal of Chemical Theory and Computation, s. 2938-2944, 2015
-
Ingår i Journal of Computational Chemistry, s. 1473-1479, 2015
-
Ingår i Journal of Molecular Structure, s. 196-202, 2015
-
Deciphering Solution Scattering Data with Experimentally Guided Molecular Dynamics Simulations
Ingår i Journal of Chemical Theory and Computation, s. 780-787, 2015
- DOI för Deciphering Solution Scattering Data with Experimentally Guided Molecular Dynamics Simulations
- Ladda ner fulltext (pdf) av Deciphering Solution Scattering Data with Experimentally Guided Molecular Dynamics Simulations
-
Force Field Benchmark of Organic Liquids. 2. Gibbs Energy of Solvation
Ingår i JOURNAL OF CHEMICAL INFORMATION AND MODELING, s. 1192-1201, 2015
-
Mobility-based prediction of hydration structures of protein surfaces
Ingår i Bioinformatics, s. 1959-1965, 2015
-
Ingår i PLOS ONE, 2015
- DOI för Deconvoluting Protein (Un)folding Structural Ensembles Using X-Ray Scattering, Nuclear Magnetic Resonance Spectroscopy and Molecular Dynamics Simulation
- Ladda ner fulltext (pdf) av Deconvoluting Protein (Un)folding Structural Ensembles Using X-Ray Scattering, Nuclear Magnetic Resonance Spectroscopy and Molecular Dynamics Simulation
-
Thermodynamics of hydronium and hydroxide surface solvation
Ingår i Chemical Science, s. 1745-1749, 2014
-
Ingår i Journal of Chemical Theory and Computation, s. 5606-5615, 2014
-
Ingår i The Journal of Physical Chemistry C, s. 7163-7173, 2014
-
CO2 and O-2 Distribution in Rubisco Suggests the Small Subunit Functions as a CO2 Reservoir
Ingår i Journal of the American Chemical Society, s. 3165-3171, 2014
-
Thiamin Function, Metabolism, Uptake, and Transport
Ingår i Biochemistry, s. 821-835, 2014
-
Ingår i Bioinformatics, s. 439-441, 2014
-
Ingår i Journal of Computational Chemistry, s. 260-269, 2014
-
Local Partition Coefficients Govern Solute Permeability of Cholesterol-Containing Membranes
Ingår i Biophysical Journal, s. 2760-2770, 2013
-
Transcription-factor binding and sliding on DNA studied using micro- and macroscopic models
Ingår i Proceedings of the National Academy of Sciences of the United States of America, s. 19796-19801, 2013
-
Ingår i Environmental Science and Technology, s. 7421-7429, 2013
-
Ingår i Peptides, s. 94-100, 2013
-
GROMACS 4.5: a high-throughput and highly parallel open source molecular simulation toolkit
Ingår i Bioinformatics, s. 845-854, 2013
-
Unexpected Effects of Cholesterol on Membrane Permeability
Ingår i Biophysical Journal, 2013
-
Ingår i Biochemistry, s. 7798-7806, 2013
-
Quantification of Solvent Contribution to the Stability of Noncovalent Complexes
Ingår i Journal of Chemical Theory and Computation, s. 4542-4551, 2013
-
Organic molecules on the surface of water droplets: an energetic perspective
Ingår i Physical Chemistry, Chemical Physics - PCCP, s. 9537-9545, 2012
-
GROMACS molecule & liquid database
Ingår i Bioinformatics, s. 752-753, 2012
-
Ingår i Journal of the American Society for Mass Spectrometry, s. 1319-1325, 2012
-
Molecular recognition in different environments: β-Cyclodextrin dimer formation in organic solvents
Ingår i Journal of Physical Chemistry B, s. 12684-12693, 2012
-
Free Energy of Separation of Structure II Clathrate Hydrate in Water and a Light Oil
Ingår i Journal of Physical Chemistry B, s. 5933-5940, 2012
-
Large Influence of Cholesterol on Solute Partitioning into Lipid Membranes
Ingår i Journal of the American Chemical Society, s. 5351-5361, 2012
-
Ingår i Journal of Chemical Theory and Computation, s. 61-74, 2012
-
Ingår i Journal of Physical Chemistry B, s. 3880-3889, 2012
-
Virus Capsid Dissolution Studied by Microsecond Molecular Dynamics Simulations
Ingår i PloS Computational Biology, 2012
- DOI för Virus Capsid Dissolution Studied by Microsecond Molecular Dynamics Simulations
- Ladda ner fulltext (pdf) av Virus Capsid Dissolution Studied by Microsecond Molecular Dynamics Simulations
-
Ingår i Journal of Chemical Theory and Computation, s. 2474-2483, 2012
-
Ingår i Physical Review E. Statistical, Nonlinear, and Soft Matter Physics, 2011
-
Ingår i Journal of Molecular Modeling, s. 3289-3297, 2011
-
Trajectory NG: portable, compressed, general molecular dynamics trajectories
Ingår i Journal of Molecular Modeling, s. 2669-2685, 2011
-
Proteins, Lipids, and Water in the Gas Phase
Ingår i Macromolecular Bioscience, s. 50-59, 2011
-
Ingår i Biophysical Journal, s. 1345-1353, 2011
-
Subunit Interface Dynamics in Hexadecameric Rubisco
Ingår i Journal of Molecular Biology, s. 1083-1098, 2011
-
On the Feasibility of Nanocrystal Imaging Using Intense and Ultrashort X-ray Pulses
Ingår i ACS Nano, s. 139-146, 2011
-
Ingår i Peptides, s. 553-559, 2011
-
Toward prediction of functional protein pockets using blind docking and pocket search algorithms
Ingår i Protein Science, s. 880-893, 2011
-
Atomistic simulation of ion solvation in water explains surface preference of halides
Ingår i Proceedings of the National Academy of Sciences of the United States of America, s. 6838-6842, 2011
-
Ingår i WIREs Computational Molecular Science, s. 710-715, 2011
-
Scrutinizing Molecular Mechanics Force Fields on the Submicrosecond Timescale with NMR Data
Ingår i Biophysical Journal, s. 647-655, 2010